-
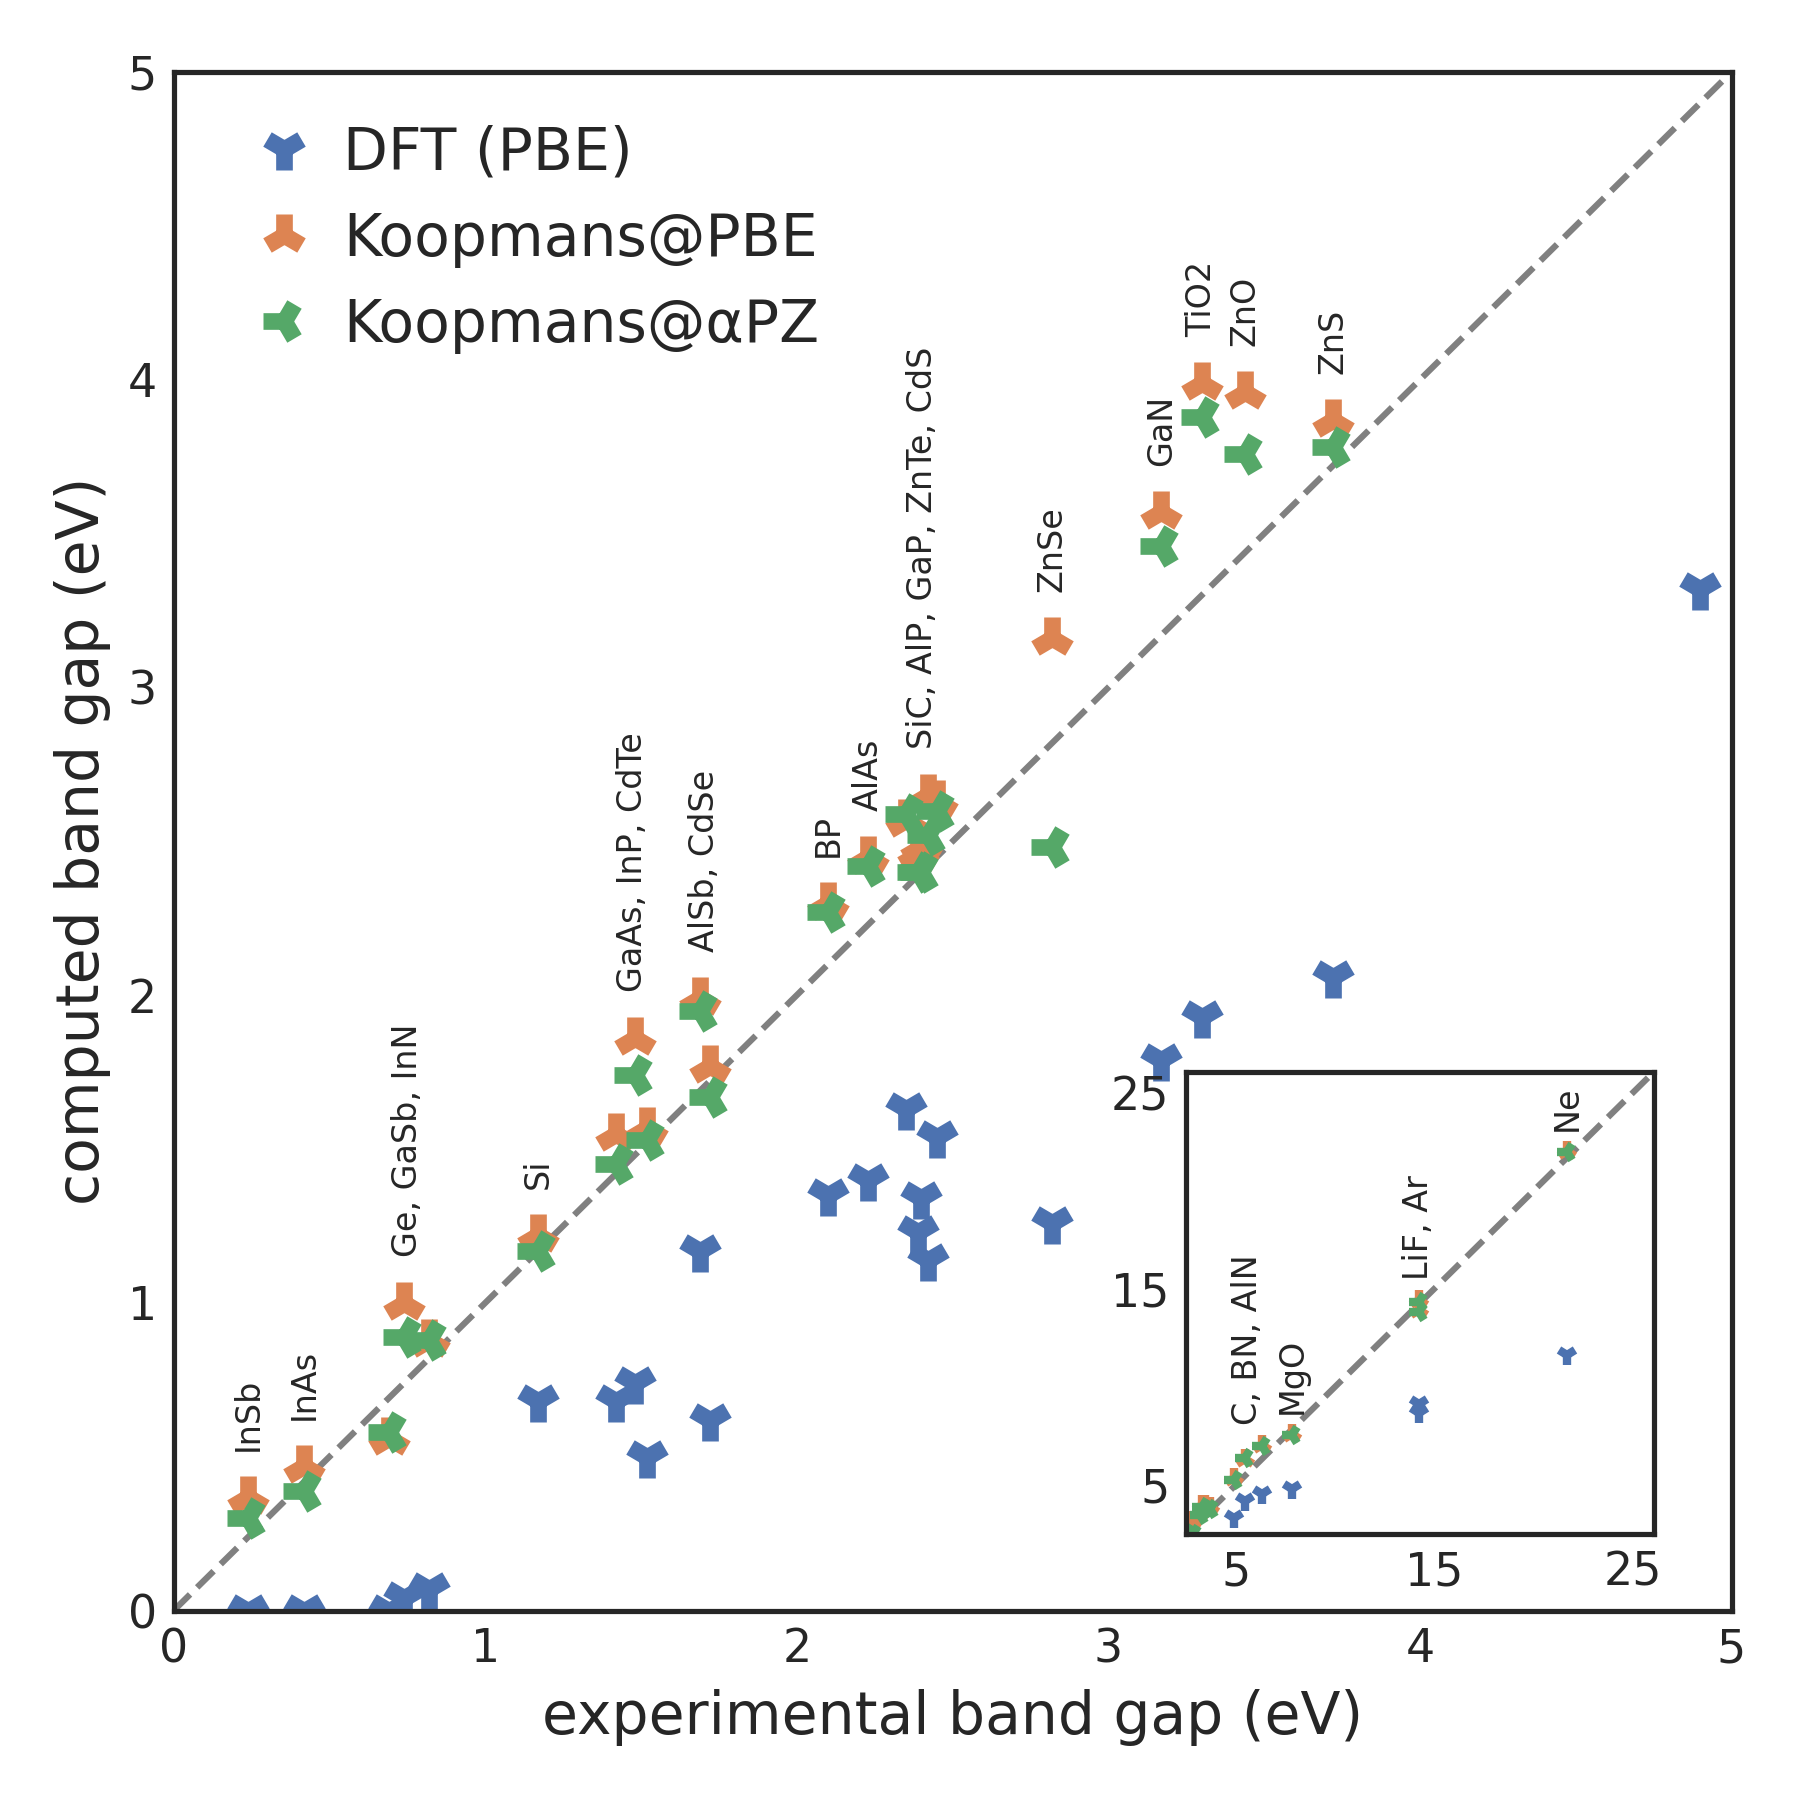
Band gaps from Koopmans functionals and DFT compared with experiment [adapted from Nguyen et al., Phys. Rev. X 2018] -
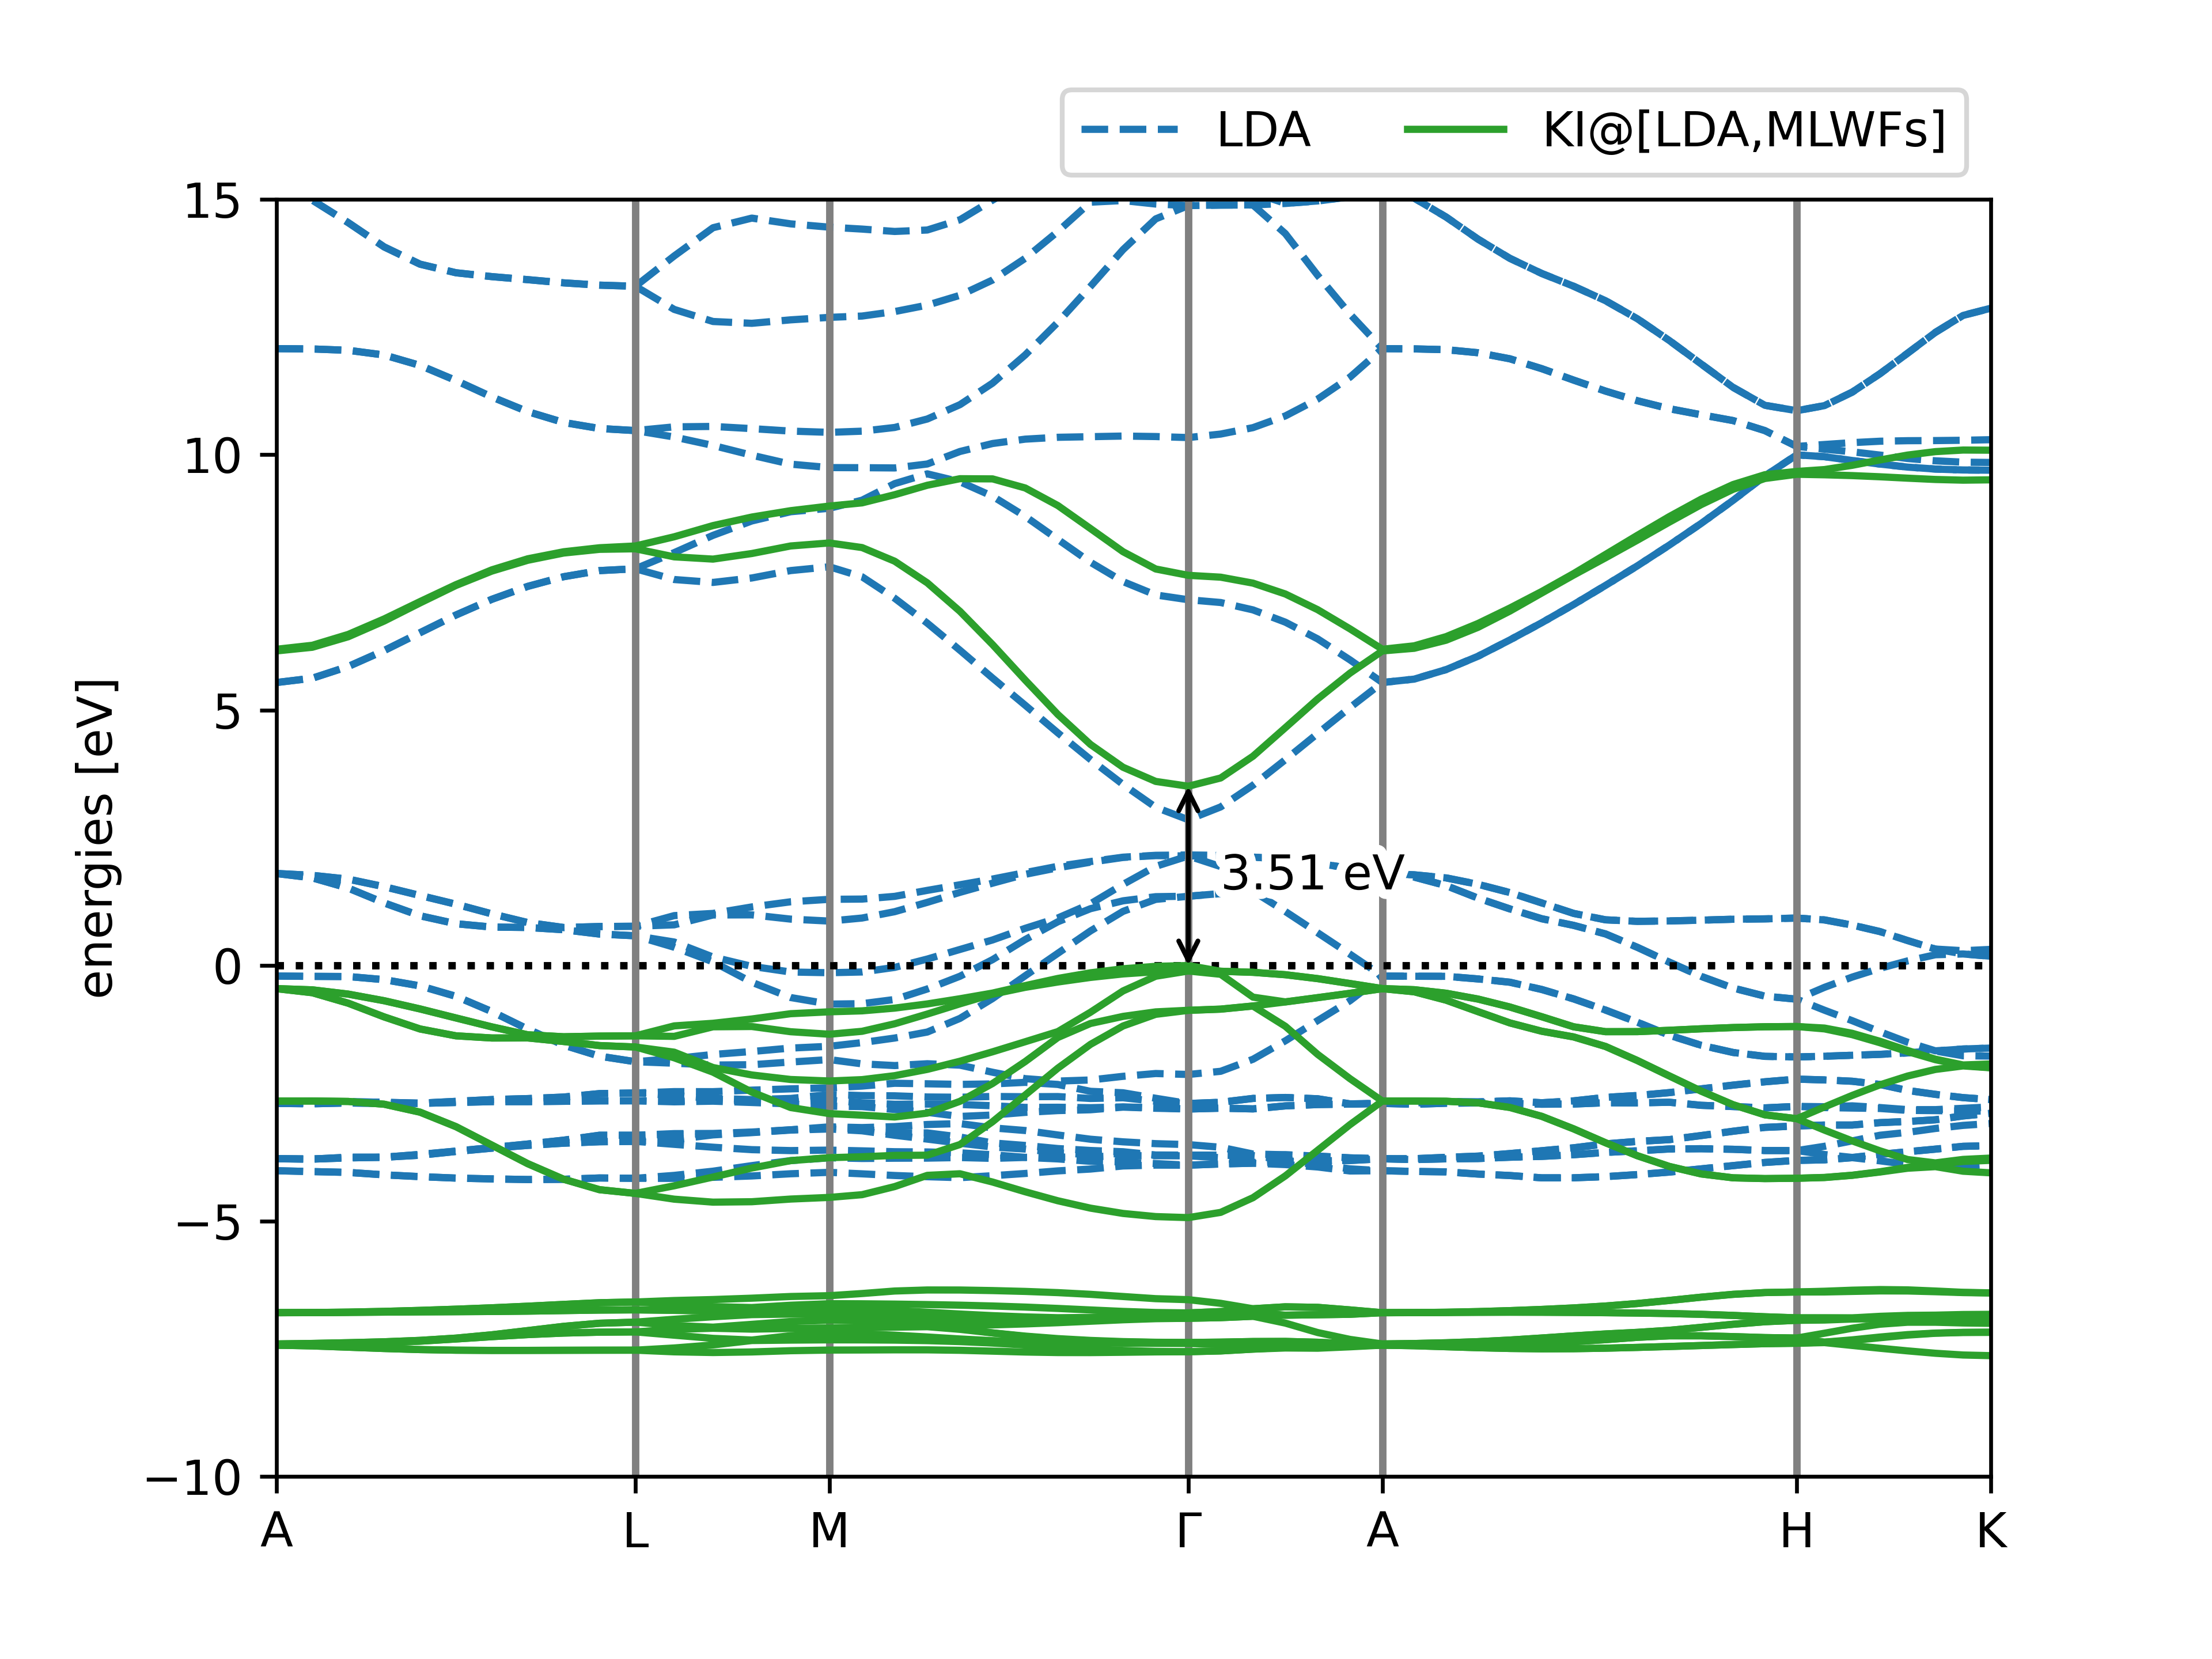
Band structure of ZnO from the KI functional, compared with LDA [Linscott et al., J. Chem. Theory Comput. 2023] -
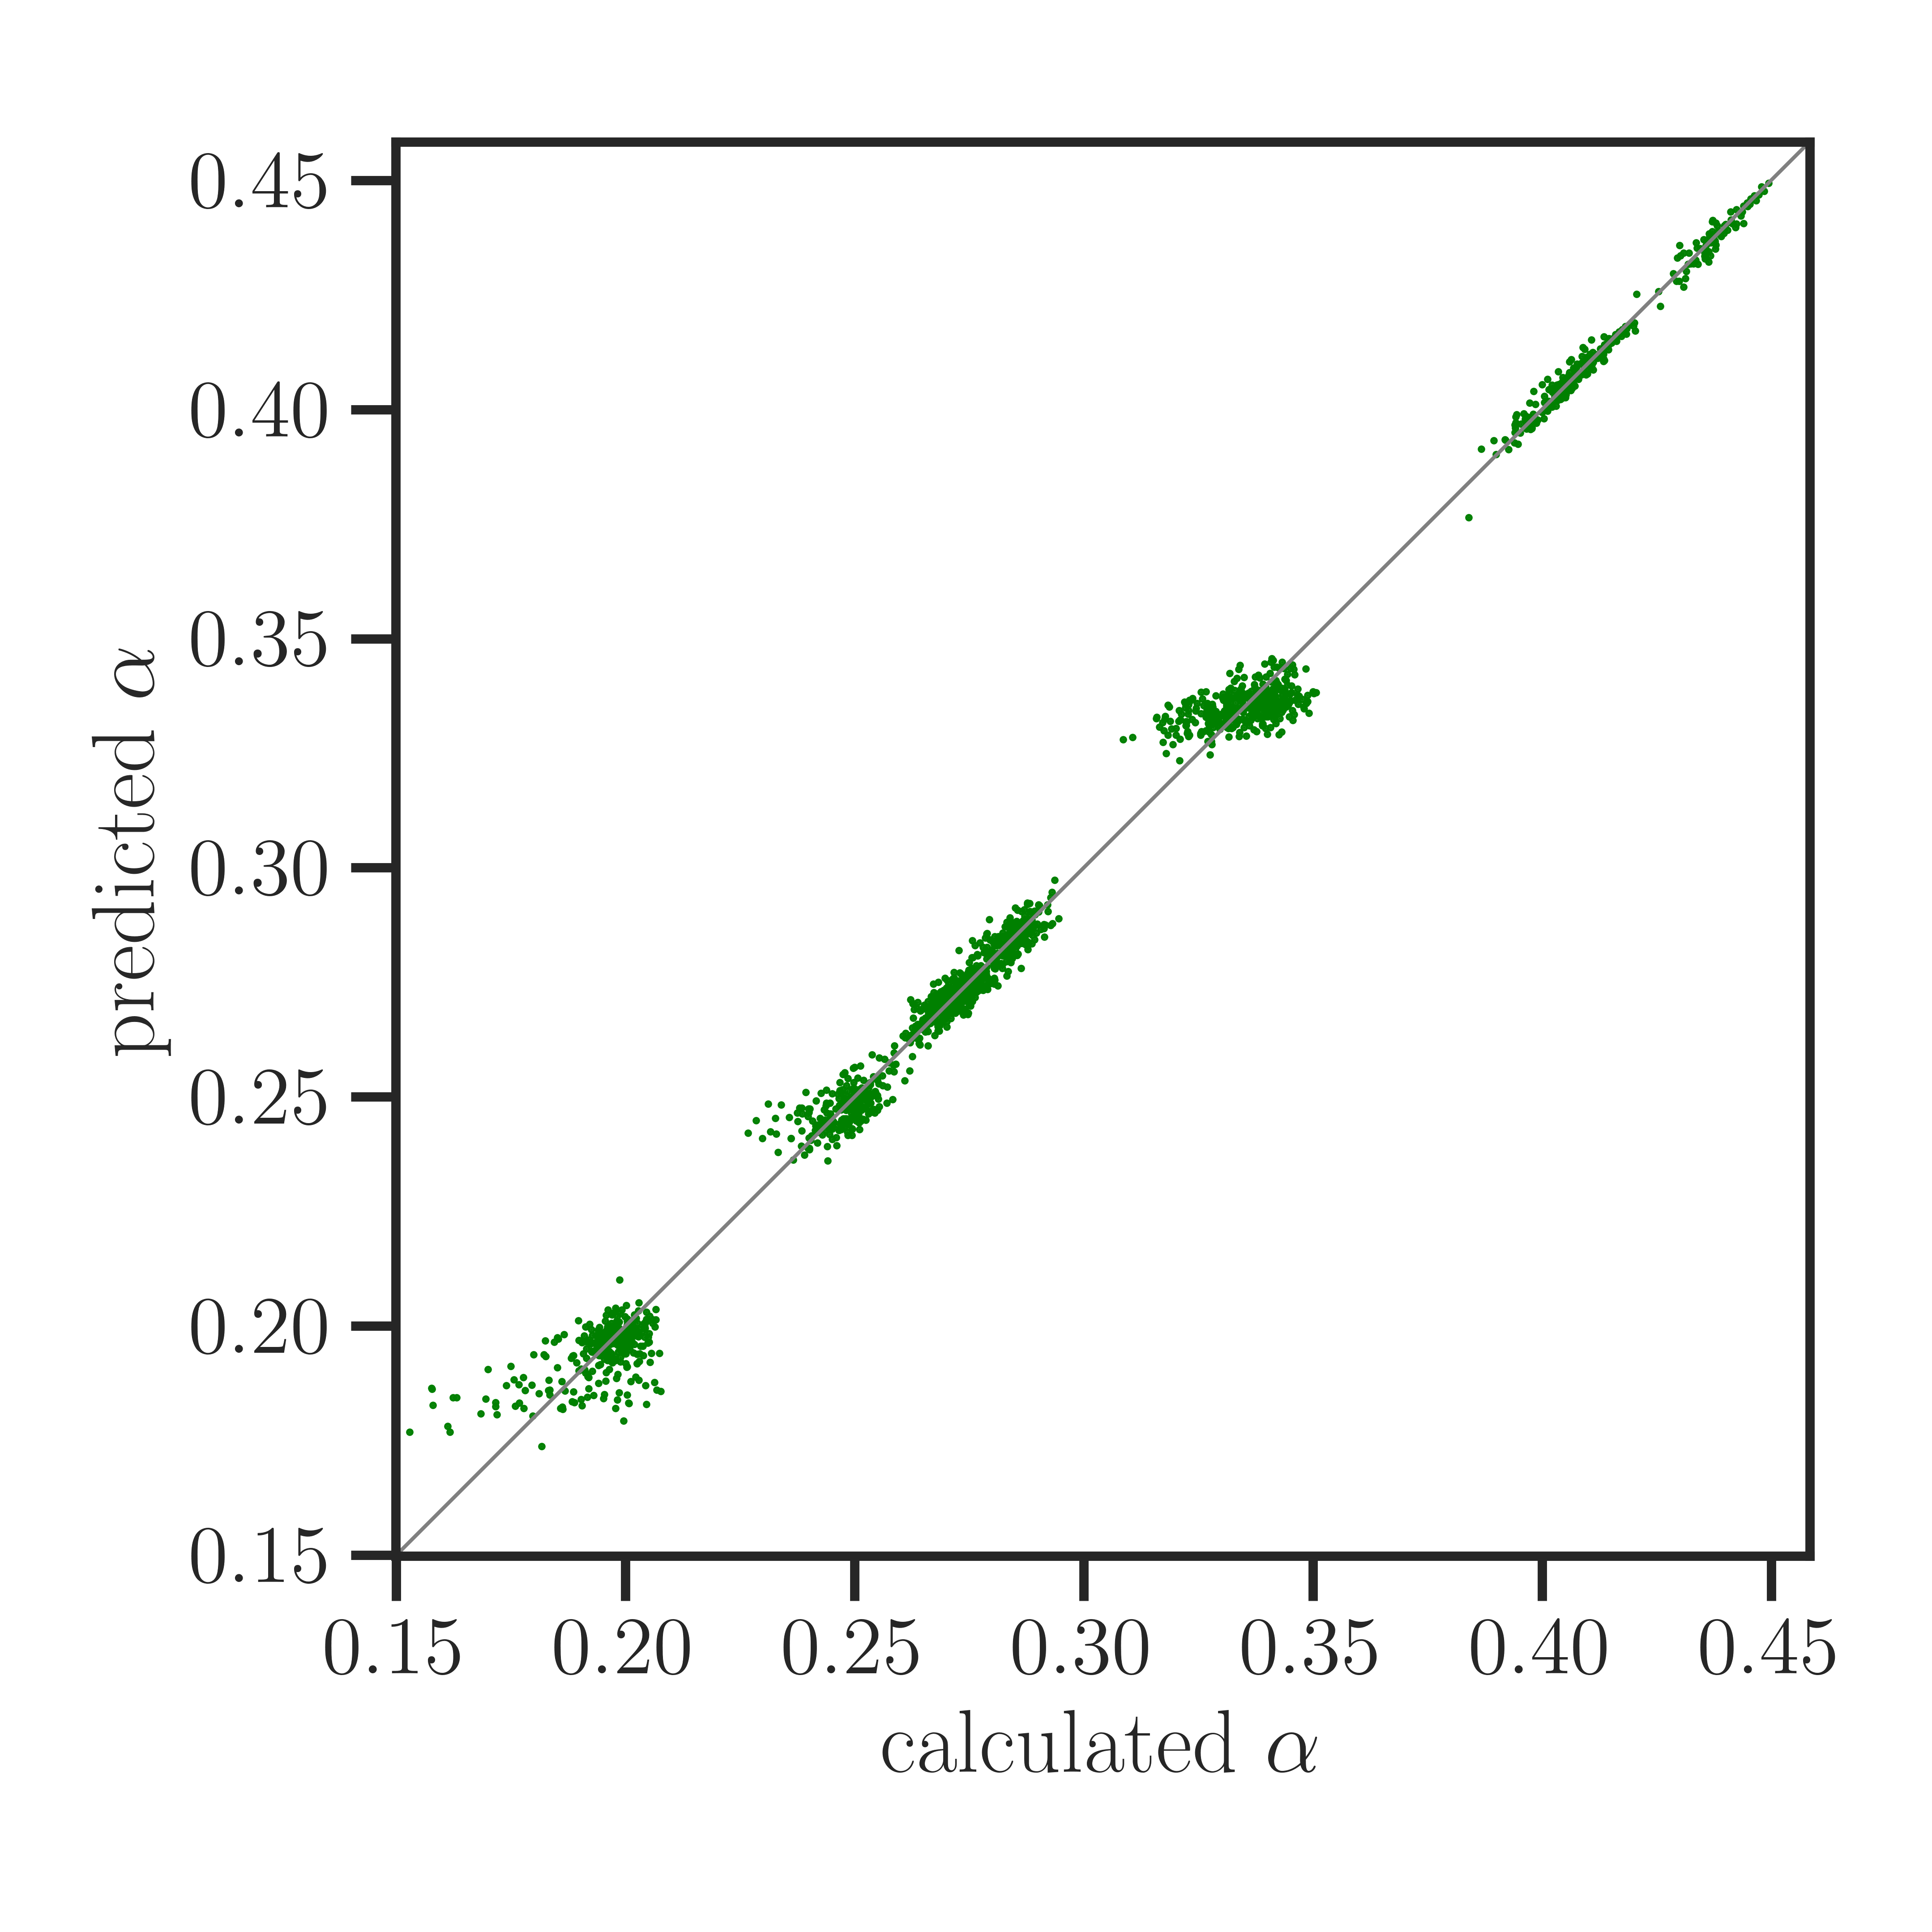
Machine-learned vs. calculated screening parameters for CsSnI3 [adapted from Schubert et al., npj Comput. Mater. 2024] -
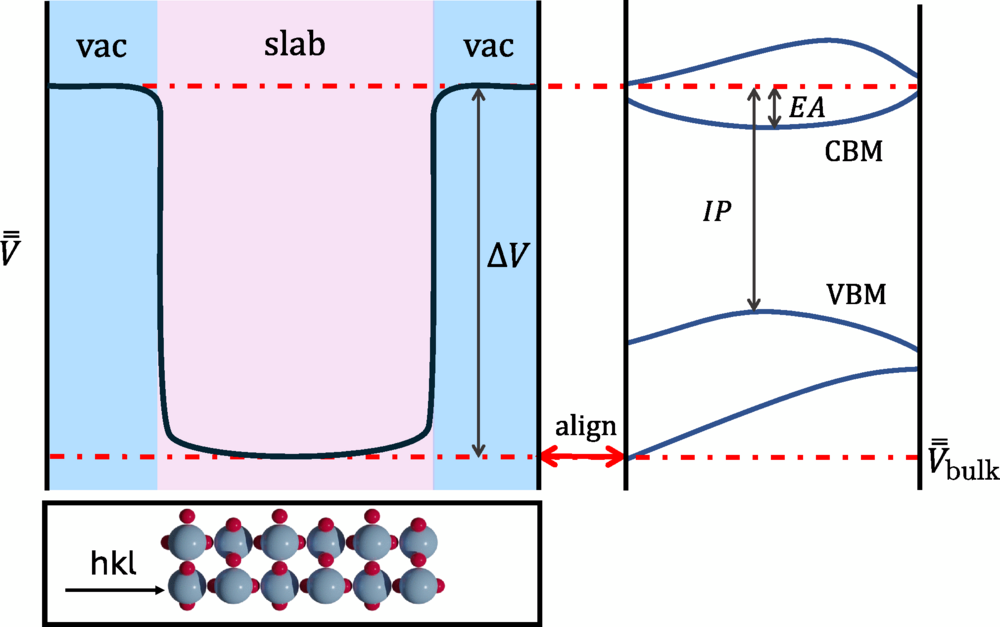
Aligning band edges to the vacuum level using a slab calculation [Stojkovic et al., Phys. Rev. Mater. 2026] -
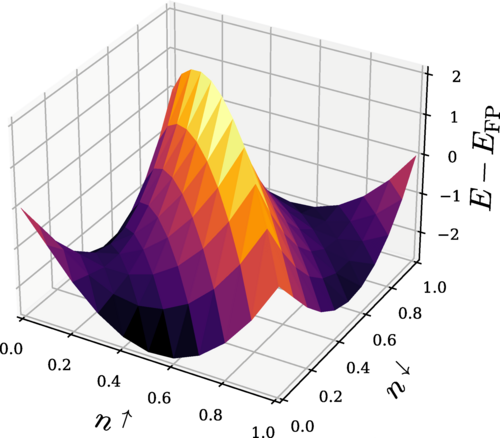
PBE energy error for He relative to the exact flat-plane condition [Burgess et al., Phys. Rev. B 2023] -
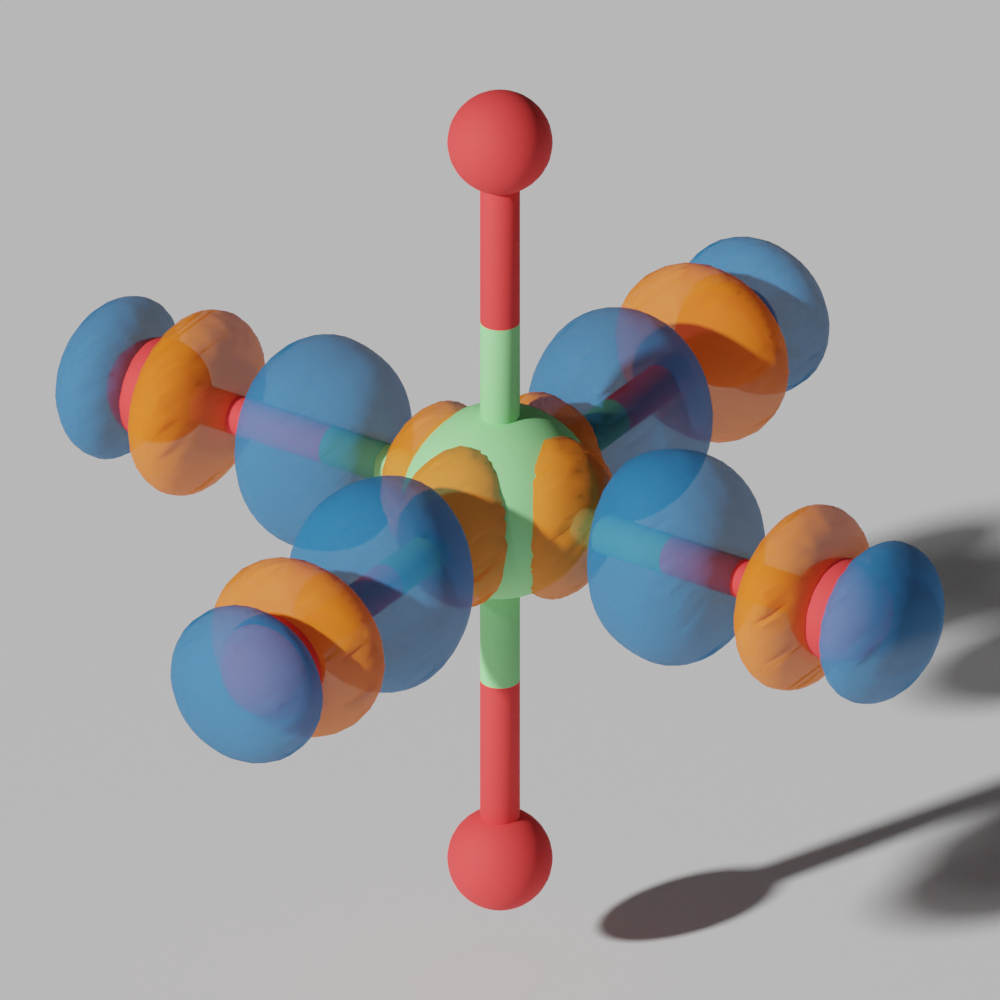
Change in effective projectors induced by inter-site terms in DFT+U+V [Linscott et al., Phys. Rev. Research 2026]
Koopmans functionals
Density functional theory (DFT) gets band gaps and photoemission spectra badly wrong because its orbital energies do not match the true energies for adding or removing an electron. Koopmans functionals fix this by imposing a generalised piecewise-linearity condition on every orbital, giving spectral properties with accuracy comparable to GW at a fraction of the cost. I lead the development of the open-source koopmans code, and work on extending these functionals, making them faster, and applying them to materials such as photocatalysts and perovskites.
Hubbard corrections and exact conditions
DFT+U is a cheap and widely used correction for materials with localised d or f electrons, but its results hinge on its parameters and its form was borrowed from the Hubbard model rather than derived from first principles. I work on computing these parameters from first principles, on understanding extensions such as DFT+U+V, and on deriving exact conditions of DFT that can guide the design of new corrections.
See papers to find out more.